
Abstract
The NF-κB signaling pathway coordinates inflammation, cell survival, and proliferation, while restraining excessive cell death to maintain immune homeostasis. Truncating mutations in RELA, encoding the NF-κB subunit p65, have been linked to autoinflammation and autoimmunity, but the underlying mechanisms remain incompletely defined. We investigated 6 patients from 5 unrelated families carrying heterozygous truncating RELA variants. Despite reduced p65 expression, patients exhibited a broad spectrum of inflammatory manifestations alongside elevated baseline and stimulus-induced proinflammatory cytokines. Functional analyses in patient-derived cells and mutant RELA-KI models showed that upstream NF-κB signaling was intact, but induction of inhibitory regulators such as IκBα and A20 was impaired. This defective feedback control shifted immune homeostasis toward amplified inflammatory responses that depended on the residual activity of the remaining functional RELA allele. Single-cell transcriptomics revealed distinct cell type–specific consequences: monocytes displayed constitutive type I IFN and NF-κB activation, B cells retained partial compensatory signaling, and T and NK cells exhibited transcriptional signatures of cell death pathways. Patient fibroblasts and mutant RELA-KI cells further confirmed enhanced TNF-induced inflammatory gene expression and hypersensitivity to apoptosis and necroptosis. These findings establish RELA haploinsufficiency as a cause of systemic immune dysregulation and link defective NF-κB feedback control to unchecked inflammation and inflammatory cell death.
Authors
Nadja Lucas, Sophia Weidler, Antonia A. Eicher, Baerbel Keller, Özlem Satirer, Adam Desrochers, Timothy J.S. Ramnarine, Mohammad Mokhtari, Sophie Elstner, Timmy Strauss, Simon W. Mages, Arek Kendirli, Oana Cristina Buzoianu, Tobias B. Haack, Lina Igel, Tim Niehues, Sandra von Hardenberg, Maria Fasshauer, Rami Abou Jamra, Hagen Ott, Ulrike Hüffmeier, Catharina Schütz, Marisa Bijwaard, Susan Wagner, Paulina Switala, Sarah Koss, Jurek Schultz, Stefanie Kretschmer, Jasmin Kümmerle-Deschner, Christine Wolf, Johanna Klughammer, Min Ae Lee-Kirsch

Abstract
Chronic kidney disease is a global health concern characterized by maladaptive repair processes that lead to kidney fibrosis. Following injury, early alterations in the extracellular matrix precede the development of kidney fibrosis and represent potential therapeutic targets to improve kidney repair. In this context, studies from our laboratory and others have shown that the matricellular protein SMOC2 can be targeted to decrease inflammation and tubulointerstitial fibrosis after kidney injury. Tubular epithelial cells (TECs), which are abundant and particularly susceptible to injury, play a central role in maladaptive repair; however, whether SMOC2 affects their function after kidney injury has not been explored. In this study, we showed that SMOC2 localized to the basement membrane of injured TECs across 3 murine models of kidney injury. Our in vitro studies demonstrate that SMOC2 induced a partial epithelial-to-mesenchymal (EMT) transition in TECs. We further demonstrated that its extracellular calcium-binding domain mediated binding to the decellularized extracellular matrix and accounted for most of its effects on TECs. Mechanistically, SMOC2 promoted partial EMT through an integrin-dependent pathway. Together, these findings provide mechanistic insight into how SMOC2 drives maladaptive repair by modulating TEC behavior and identify its calcium-binding domain as a key functional mediator.
Authors
Schrodinger Cenatus, Peng Gao, Nathalie Henley, Caroline Lamarche, Xue-Song Liu, Frédérick A. Mallette, Jonatan Barrera-Chimal, Casimiro Gerarduzzi

Abstract
Systemic lupus erythematosus (SLE) is a progressive autoimmune disease that affects multiple organs and tissues, with lupus nephritis (LN) as one of its most severe complications. Although LN progression is associated with compromised permeability of human renal glomerular endothelial cells (HRGECs), the underlying mechanisms are not fully defined. Herein, we demonstrate that aberrant glycolysis drives this glomerular endothelial barrier defect by suppressing the transcription of tight junction (TJ) genes. Mechanistically, circulating self-DNA in SLE plasma acts as a ligand that activates the cGAS/STING pathway in HRGECs, driving aberrant glycolytic adaption. The resulting glycolytic product, lactate, serves as a substrate for protein lactylation, leading to extensive lactylation and subsequent ubiquitination of enhancer of zeste homolog 2 (EZH2). In consequence, EZH2 deficiency results in reduced H3K27me3 levels, thereby suppressing the transcription of TJ genes. In a self-DNA–induced SLE model, inhibition of cGAS/STING signaling or lactate production effectively restored the integrity of TJs of HRGECs and concurrently alleviated key LN symptoms. Together, lactate programs lactylation and ubiquitination of EZH2 to impair the glomerular endothelial barrier in human SLE.
Authors
Jiaxin Lei, Xingyu Zhai, Yixin Wang, Ying Li, Lei Li, Mengdi Liu, Jing Guo, Lingyi Li, Zhezhuyun Chen, Qinghua Cao, Zhichun Liu, Ting Liu, Lin Xu, Zhenke Wen

Abstract
Bone marrow–derived circulating monocytes continuously replenish intestinal macrophages, which become dysregulated in inflammatory bowel disease (IBD) and contribute to disease pathology. The origins of this dysregulation remain poorly understood. Here, we investigate the reprogramming of circulating monocytes in IBD prior to tissue recruitment using single-cell transcriptomic, epigenomic, and functional approaches. We characterize blood monocyte heterogeneity in newly diagnosed, treatment-naive IBD patients and healthy controls and show that monocytes in Crohn’s disease (CD) display a distinct transcriptional profile and altered distributions across inferred developmental trajectories; less pronounced changes were observed in ulcerative colitis (UC). We link CD-associated transcriptional changes to alterations in chromatin accessibility and identify NF-κB, EGR, KLF, and AP-1 family transcription factors as putative regulators of an inflammatory gene program in blood monocytes from patients with CD. We uncover a potential role for IFN-γ in priming blood monocytes for inflammatory function in CD by limiting their capacity to be regulated by IL-10. Finally, we show that the transcriptional and functional alterations in monocytes from patients with CD are maintained in monocyte-derived cells from the intestine. Together these data suggest that intestinal macrophage dysfunction in CD is, at least in part, pre-established by systemic signals prior to tissue recruitment.
Authors
Eve Hornsby, Radha Gadhok, Inva Hoti, Eva Wozniak, James R. Boot, Emma Connick, Paul Stevens, Holly Creed, Amy Lewis, Andrew Silver, James O. Lindsay, Andrew J. Stagg
Abstract
The mechanisms by which e-cigarette vaping (EV) affects lung health remain unclear. Clusters of EV-associated lung injury indicate that EV damages distal lung parenchyma and increases vulnerability to second-hit injury, including respiratory viral infections. Using human lung endothelial and epithelial cells and precision-cut lung slices, we investigated the mechanisms underlying distal lung cell injury and repair triggered by brief (24-hour) EV exposure. RNA-seq of lung tissue from golden Syrian hamsters evaluated the persistence of lung stress responses (10 days after 5 days of EV exposure) and the effect of EV on host defense against influenza A virus and SARS-CoV-2. EV disrupted the barrier function of human distal lung cells through JNK stress response signaling, triggered autophagy with impaired flux, suppressed mTOR signaling and cell proliferation, and culminated in apoptosis. Transcriptional responses in EV-exposed hamster lungs revealed persistent activation of JNK signaling, autophagy, barrier dysfunction, tissue remodeling, and impaired Th1 immunity. EV increased SARS-CoV-2 viral burden, downregulated antiviral genes (Ifit1, Isg15, Nfkbia), and amplified oxidative stress and IL-12 signaling. Short-term EV exposure triggered stress-induced distal lung cell injury with persistent changes in antiviral immunity and molecular pathways associated with tissue remodeling. That may increase susceptibility to viral infections and contribute to lung disease.
Authors
Tanner C. Rivera, Kelly S. Schweitzer, Christina Cornell, Jordan Nall, Nicholas Egersdorf, Courtney Moeder, Riley A. Cooney, Eszter K. Vladar, Steve D. Groshong, Gregory P. Downey, James P. Bridges, Richard Bowen, Hong Wei Chu, Irina Petrache

Abstract
Lineage plasticity, or transdifferentiation, is increasingly recognized as a resistance mechanism to androgen receptor (AR) inhibition in prostate cancer. Lineage plasticity is characterized by loss of AR signaling and epithelial differentiation, along with activation of stemness-associated pathways, epithelial-mesenchymal transition, or alternative differentiation programs such as neuroendocrine prostate cancer (NEPC). Loss of the tumor suppressors TP53 and RB1 is common in tumors exhibiting lineage plasticity; however, the mechanisms by which TP53/RB1 loss promotes this phenotype remain poorly understood, and effective treatments are limited. Using multiomic profiling of TP53/RB1-loss prostate cancer models, we identified alterations in chromatin accessibility, DNA methylation, and gene expression associated with lineage plasticity. Importantly, many pathways activated upon TP53/RB1 loss could be blocked through BET bromodomain inhibition. TP53/RB1-deficient cells also harbored widespread DNA methylation changes that silenced pathways linked with restraining lineage plasticity. Combined BET bromodomain and DNA methyltransferase (DNMT) inhibition was more effective than single-agent treatment in suppressing growth of TP53/RB1-loss models exhibiting a stem-like or NEPC program. This was partly explained by abrogation of discrete lineage plasticity pathways modulated by each agent. Altogether, our work suggests combined BET bromodomain and DNMT inhibition is a promising therapeutic approach for prostate tumors exhibiting lineage plasticity.
Authors
William K. Storck, Diana Flores, Anbarasu Kumaraswamy, Zhi Duan, Shrabastee Chakraborty, Chao Zhang, Eva Rodansky, Dhruv Khokhani, Olivia A. Swaim, Karan Bedi, Raymond G. Cavalcante, Canping Chen, Faming Zhao, Ya-Mei Hu, Zheng Xia, Ryan J. Rebernick, Marcin Cieslik, Rahul Mannan, Somnath Mahapatra, Arul M. Chinnaiyan, Aaron M. Udager, Joshua A. Kuleape, Catherine R. Alumkal, Hannah N. Beck, Peter S. Nelson, Colm Morrissey, Michael C. Haffner, Leigh Ellis, Yuzhuo Wang, Joel A. Yates, Joshi J. Alumkal

Abstract
Plasmodium falciparum sporozoite (PfSPZ) vaccines, composed of aseptic, purified, live parasites that arrest during or just after liver-stage development, show excellent safety and efficacy in humans. They can induce complete protection against Pf infection, mediated primarily by cellular immune responses against parasite antigens expressed in hepatocytes. Current PfSPZ vaccines rely on the West African PfNF54 parasite, which uniquely produces high numbers of PfSPZ in mosquitoes, facilitating manufacturing efficiency. However, PfNF54 has relatively low hepatocyte infectivity, limiting potency. We created hybrid pan-African Pf strains by genetically crossing PfNF54 with East African Pf strains. The hybrid, AV27, was selected for development based on balanced contribution of parental genomes, high PfSPZ production, and high liver-stage infectivity. As compared with NF54-based PfSPZ vaccines, we expect AV27-based vaccines will have greater and broader efficacy at lower doses due to higher liver-stage infectivity and inclusion of unique East African CD8+ T cell epitopes.
Authors
Lucia Pazzagli, Bethany Jenkins, Ankit Dwivedi, Asha Patil, Yonas Abebe, Tales V. Pascini, Urvashi Rai, Priya Gupta, Nastaran Rezakhani, Chakshu Gandhi, Yiwei Yang, Sudhir Kumar, Mohd Kamil, Gigliola Zanghí, Manuel Llinás, Stephen L. Hoffman, Joana C. Silva, Ashley M. Vaughan, B. Kim Lee Sim
Abstract
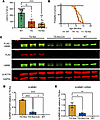
Autosomal dominant leukodystrophy (ADLD) is a fatal adult-onset CNS demyelinating disorder for which no treatment exists. The majority of ADLD cases are caused by duplications of the lamin B1 (LMNB1) gene, resulting in increased LMNB1 expression. While reducing LMNB1 levels represents a logical therapeutic strategy, its efficacy has not been previously demonstrated in any in vivo model. Mouse models with oligodendrocyte-specific human LMNB1 (hLMNB1) overexpression recapitulate salient features of ADLD. Using a modified version of this model, where hLMNB1 can be inducibly downregulated, we demonstrated that hLMNB1 reduction can prevent or substantially ameliorate disease progression. Therapeutic effects were maximized when hLMNB1 reduction was induced before expected symptom onset, resulting in improvements in behavioral, biochemical, histopathological, and survival measures relative to those of untreated animals. Reducing hLMNB1 levels after symptom onset led to improved survival but mixed results for other disease phenotypes. In addition, we identified potential biomarkers that track disease progression. Furthermore, we demonstrated that near-complete knockdown of murine LMNB1 expression in adulthood did not result in any overt CNS phenotype. Together, these results provide a proof of concept supporting LMNB1 reduction as a therapeutic strategy and offer a rationale for treatments aimed at lowering levels of this protein in ADLD.
Authors
Nathan Herdman, Kaveh Moradi, Bruce Nmezi, Anushe Munir, Krizchelle A. Magtoto, Fang Liu, Mara Sullivan, Xuemei Zeng, Thomas K. Karikari, Quasar S. Padiath

Abstract
Autoimmune Addison’s disease (AD) is a rare but life-threatening disorder caused by immune-mediated destruction of the adrenal cortex, and progress in therapy has been limited by insufficient mechanistic insight. Here, we establish a model of experimental autoimmune adrenalitis that recapitulates key features of AD and reveals sex-dependent differences in disease manifestation within the model. Immunization with peptides derived from the adrenal self-antigen CYP11A1 induces corticosterone insufficiency. We show that autoimmune adrenalitis is driven by IFN-γ produced by self-reactive CD4+ T cells, promoting granulomatous inflammation in the adrenal cortex. Together, these findings identify IFN-γ as a central effector of autoimmune adrenalitis and suggest that targeting the IFN-γ pathway may represent a potential therapeutic strategy for AD.
Authors
Arina Andreyeva, Juraj Michalik, Veronika Niederlova, Veronika Cimermanova, Ales Drobek, Radislav Sedlacek, Jan Prochazka, Juraj Labaj, Olha Fedosieieva, Waldemar Kanczkowski, Peter Draber, André Sulen, Ondrej Stepanek, Aleš Neuwirth

Abstract
Right ventricular pressure overload (RVPO) is a critical pathophysiological feature of numerous pediatric cardiovascular diseases. Transverse tubules (T-tubules) form the foundation for efficient excitation-contraction coupling in mature cardiomyocytes. We hypothesized that RVPO impairs T-tubule maturation through the regulatory protein bridging integrator 1 (BIN1). In right ventricular samples from children with tetralogy of Fallot, characterized by RVPO, and in a neonatal rat RVPO model induced by pulmonary artery banding (PAB), T-tubule maturation was disrupted. RNA-seq revealed significant downregulation of T-tubule–associated genes, with Bin1 among the most suppressed. Bin1 overexpression restored T-tubule maturation in PAB rats. ATAC-seq showed reduced chromatin accessibility at Bin1 loci; motif analysis identified Mef2d (myocyte enhancer factor 2D) as the top enriched transcription factor. Mef2d knockdown rescued Bin1 expression and T-tubule maturation, and mutation of the Mef2d binding sites within the Bin1 promoter abolished the inhibitory effect of Mef2d on Bin1 promoter activity. This study delineates a phenomenon and a mechanism of cardiomyocyte maturation under pathological stress. The findings not only advance our understanding of this most pivotal event in postnatal cardiac development but also unveil a potential therapeutic direction for pediatric cardiovascular diseases associated with RVPO.
Authors
Yuqing Hu, Yiting Xue, Xudong Chen, Linghui Kong, Debao Li, Zheng Wang, Sixie Zheng, Siqi She, Hao Li, Sijuan Sun, Hao Chen, Lijun Chen, Peisen Ruan, Kai Wang, Lincai Ye
No posts were found with this tag.


